Cytiva Partners with Mycenax Biotech to Build AI-Driven and Future-Ready Downstream Bioprocessing
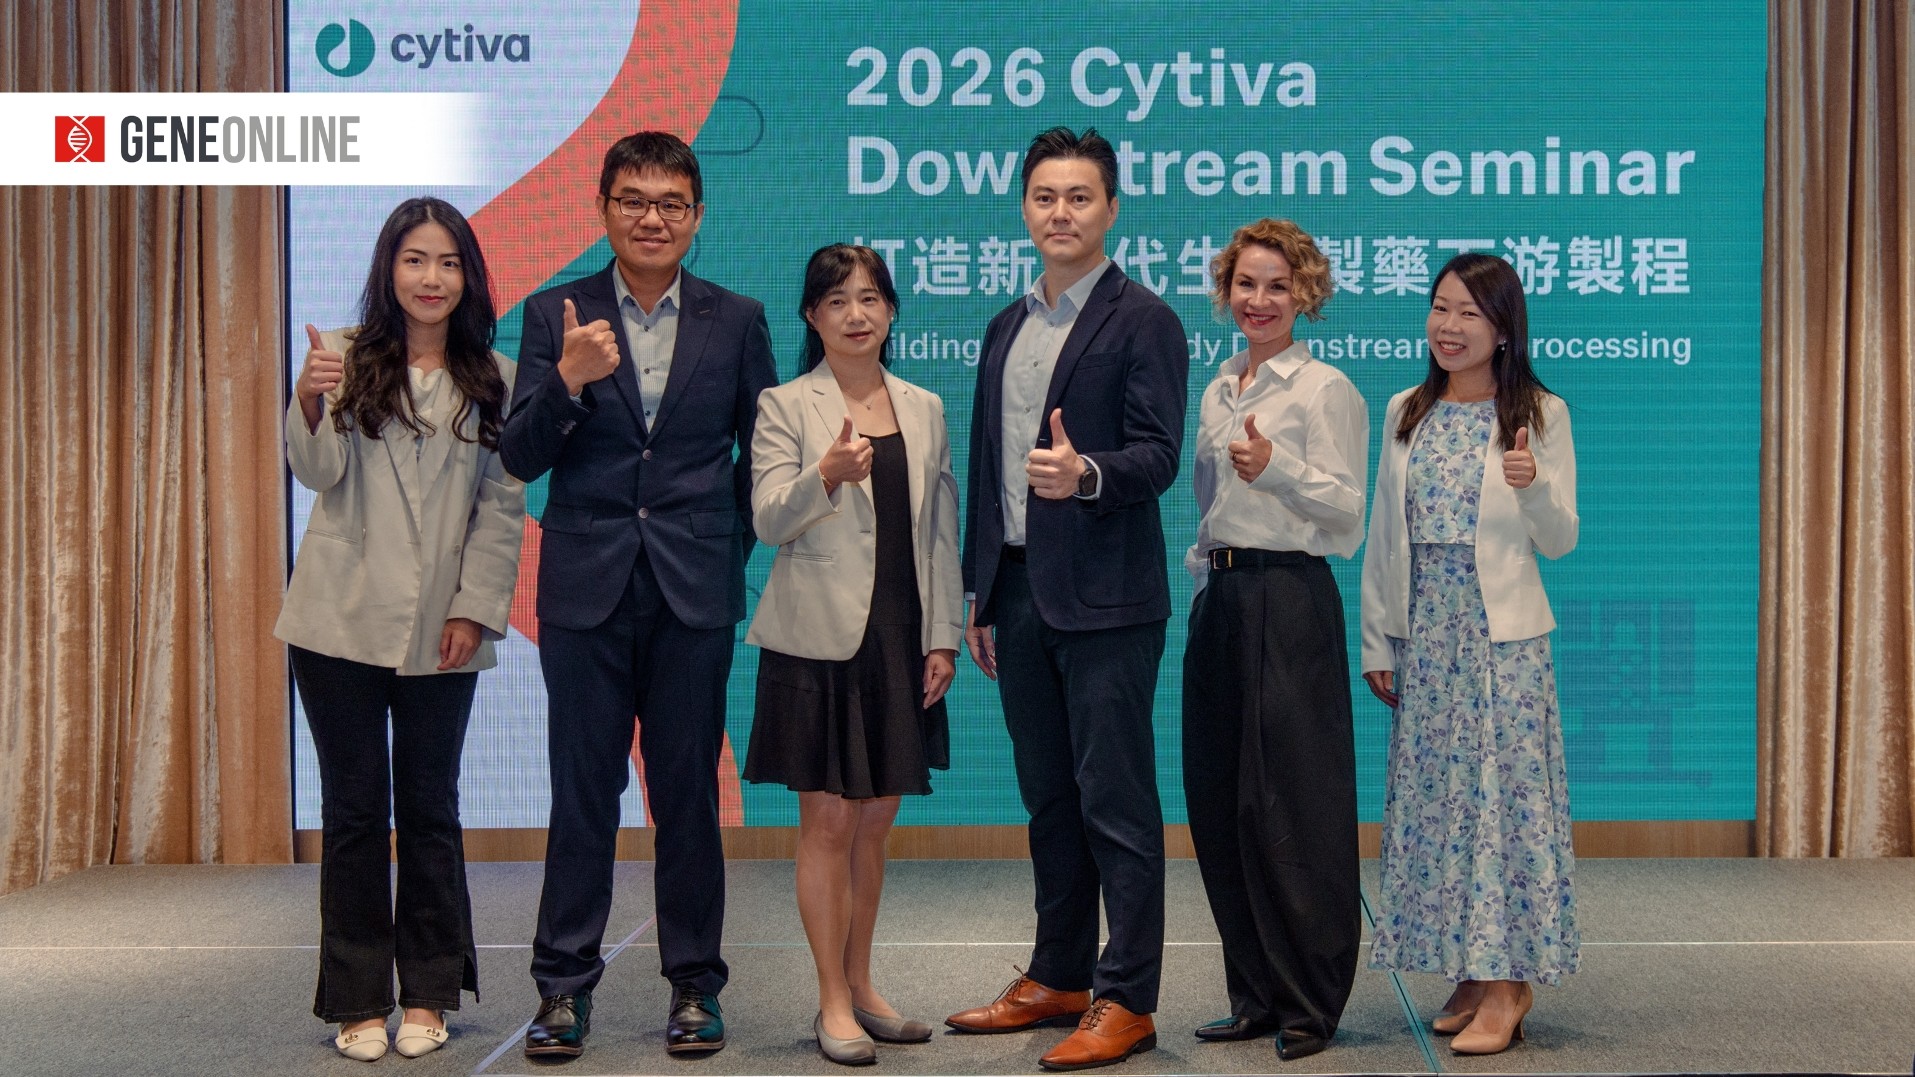
As the global biopharmaceutical industry evolves rapidly, the downstream processes for high-concentration biologics and emerging therapies are becoming increasingly complex. Balancing production flexibility, regulatory compliance, and cost control has thus become the key to commercial success for biotech firms and contract development and manufacturing organizations (CDMOs). To address these challenges, Cytiva, world’s leading life sciences equipment and service provider, organized a seminar titled “Building the Future-Ready Downstream Bioprocessing” in Taipei on July 3, 2026. The event gathered over 100 experts in process engineering and regulatory affairs to explore GMP-compliant ways for boosting manufacturing efficiency and capacity through the deployment of continuous purification, single-use technology platforms, and AI-driven digital twin solutions.
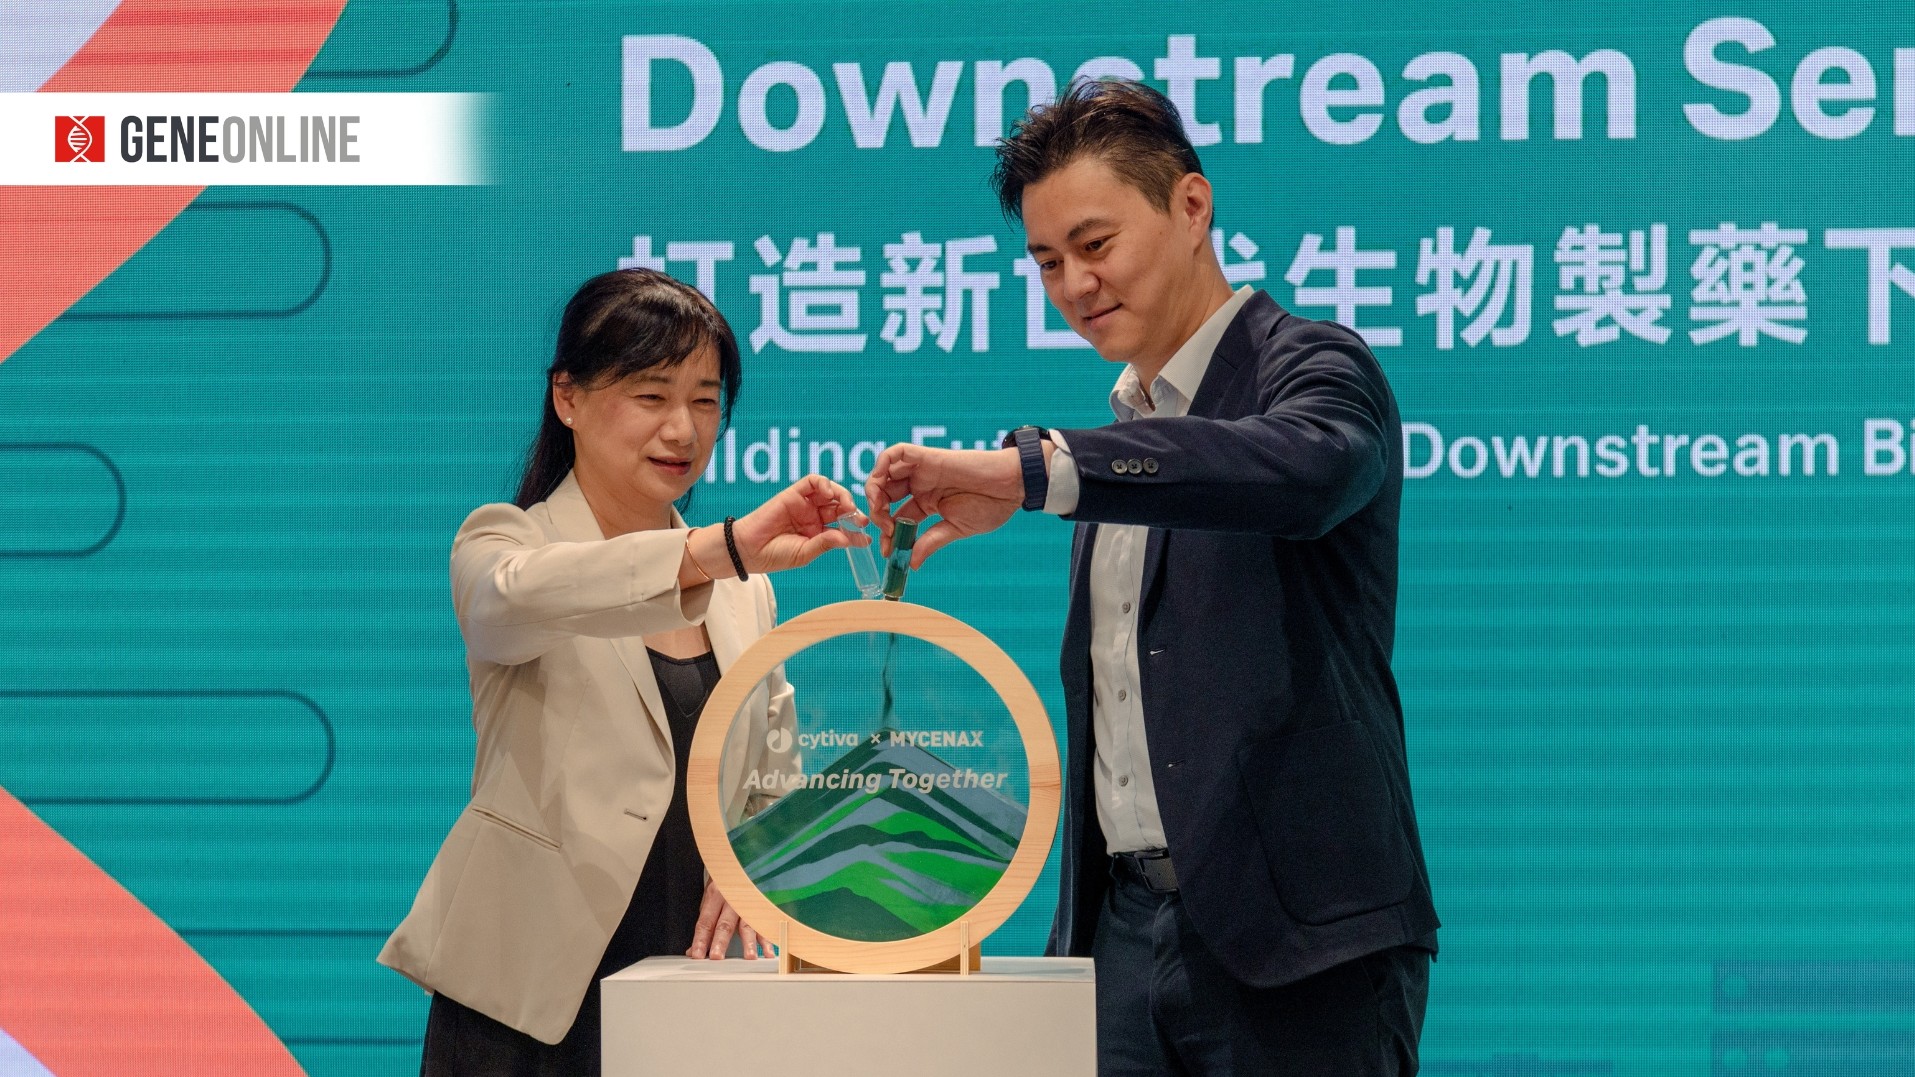
The event commenced with a symbolic sand art creation ceremony featuring Jeff Wang, Cytiva’s Business Head of Taiwan and Dr. Pei-Jiun Chen, CEO and President of Mycenax Biotech. Titled “Advancing Together,” the artwork centers on the concept of “integration,” not only celebrating the steadfast collaboration and mutual trust between the two companies over the past two decades, but also declaring their ongoing commitment to accelerating the transformation of Taiwan’s biopharma sector.
In addition to a rich lineup of expert presentations, GeneOnline exclusively interviewed these two industry leaders on-site. During the interview, they discussed the strategic partnership between Cytiva and Mycenax and the future outlook for Taiwan’s CDMO sector from diverse perspectives, including technological exchange, industry trends, and international markets.
Optimizing Downstream Bioprocesses with GoSilico AI
In the opening session of the seminar, Dr. Pei-Jiun Chen, CEO and President of Mycenax, analyzed the key challenges in downstream bioprocessing from the big-picture perspective of a one-stop CDMO, using real-world case studies to illustrate how Mycenax has overcome these challenges by adopting AI tools. She began by highlighting the highly competitive and time-sensitive nature of modern life sciences and pharma industries, noting that virtually all companies are racing to advance their drug candidates into clinical trials as quickly as possible to capitalize on market opportunities. However, with the rise of emerging modalities such as bispecific (multispecific) antibodies and antibody-drug conjugates (ADCs), the complexity of process development scales exponentially, making it harder to shorten development timelines.
For example, downstream chromatographic purification involves multiple interdependent variables such as pH value, flow rate, buffer salt concentration, and column size. This leaves R&D personnel struggling to balance purity and product recovery. Relying on traditional trial-and-error experimental designs is time-consuming, risks wasting valuable proteins, and drives up development costs.
To resolve this pain point, Mycenax has taken the lead in Taiwan’s industry by pioneering the adoption of Cytiva’s GoSilico™ AI platform. This digital twin solution is specifically designed for the development and optimization of column chromatography processes. At its core, it uses experimental data to build mechanistic models, which perform computer simulations based on known physical and chemical phenomena in chromatography and incorporate AI to optimize the process. Based on real-world experimental data, the system comprehends the interactions between proteins and resins under various conditions. Upon data entry, it generates a highly realistic digital twin, accurately simulating and predicting chromatographic profiles, recovery rates, and purity.
In her presentation, Dr. Chen shared a case study involving a client with an antibody candidate that had entered human clinical trials. While their original chromatography process could yield antibodies of extremely high purity, the recovery rate was less than 60%, which could lead to a surge in production costs when scaling up to GMP-compliant commercial production, thus undermining the product’s market competitiveness. Facing tight deadlines, however, the client could not afford the material and time costs associated with large-scale experimentation. In response, the Mycenax team introduced the GoSilico system to optimize the process by leveraging AI and digital twin to tackle efficiency bottlenecks in the existing purification steps.
Operationally, the team imported data from five sets of real-world chromatography experiments into the GoSilico system. Following a two-week calibration by Cytiva’s experts, a highly realistic digital twin model was created. The model demonstrated exceptional performance by executing over 3,000 simulations in a single day, calculating antibody recovery rates and purity under various combinations of buffer formulations, pH values, and washing volumes to successfully identify the optimal balance between throughput and quality.
Using 20 target conditions selected for validation, the team tested the model’s accuracy, achieving less than 5% error in predicted purity and less than 10% error in recovery rates. During final process validation, the discrepancy between actual data and model predictions was a mere 1%. By adopting this AI-selected set of parameters, the new process reduced the number of washing and elution steps from four to two, streamlining the GMP commercial production process and boosting product recovery by 11%.
According to Dr. Chen, the GoSilico system shortens timelines, reduces raw material consumption, and aligns with global regulatory trends such as the Quality by Design (QbD) promoted by the FDA. Integrating AI simulation data into regulatory dossiers can demonstrate to authorities a comprehensive understanding of both process robustness and the optimal operational ranges required for safety and quality. This evidence has the potential to expedite the review timelines for new drug applications. “By leveraging AI and digital twins, Mycenax can ensure that the developed processes are fully compatible with large-scale GMP production, meeting clients’ expectations for high-quality, high-throughput drug manufacturing and commercialization,” she added.
Dr. Chen also noted that AI is transforming bioprocessing from an experience-driven approach to a data-driven one. Due to constraints in manpower, resources and time, R&D teams cannot conduct thousands of physical experiments, but AI systems can complete virtual experiments in a single day. This human-machine collaboration broadens scientists’ horizons and stabilizes drug development, delivering real value to clients and patients instead of replacing humans.
Building a Profitable Business Model Based on Mutual Trust
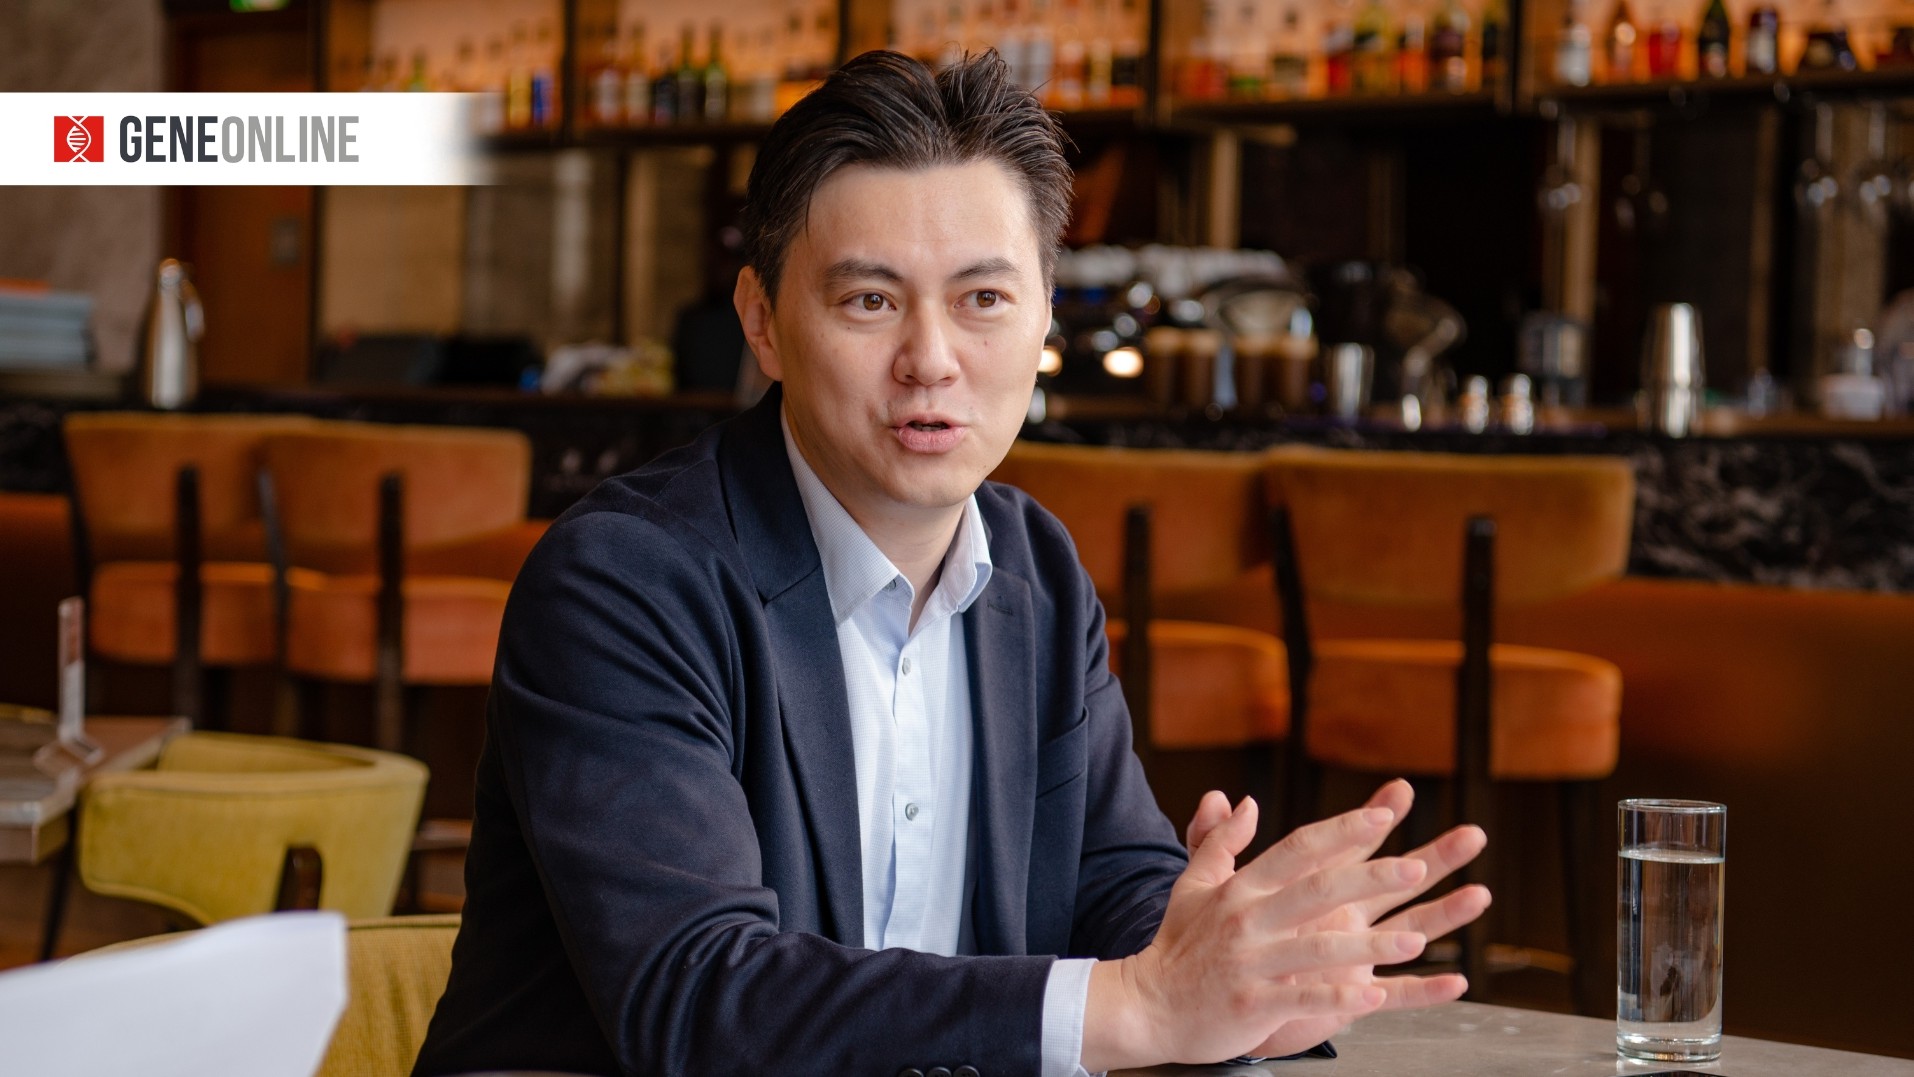
The successful implementation of the GoSilico system represents more than just a technological breakthrough. It is also a testament to the long-standing strategic partnership between Cytiva and Mycenax. Following the seminar, GeneOnline conducted exclusive interviews with Jeff Wang, Cytiva’s Business Head of Taiwan, and Dr. Pei-Jiun Chen, Mycenax’s CEO and President. Starting from the catalyst of this digital transformation, the two industry leaders offered insights into how Taiwanese CDMOs can find their niche and chart their future course amid intensifying global biotech competition.
Wang first recalled that GoSilico is not the first collaboration between Cytiva and Mycenax. Over the past two decades, Mycenax has not only been among the first in Taiwan to integrate Cytiva’s single-use technology into GMP production lines but has also led the industry in adopting continuous processing. From an equipment supplier’s perspective, he highlighted that within Taiwan’s conservative life sciences sector, Mycenax’s openness to innovation makes it Cytiva’s ideal partner for validating new concepts locally.
Both executives shared a clear consensus regarding the motivation for adopting the GoSilico platform: AI should not be embraced simply for its own sake. Wang observed that although the industry has frequently heralded “the dawn of the AI era” in recent years, clients primarily care about whether new tools can deliver tangible benefits. Therefore, merely employing AI to analyze data and generate reports is clearly inadequate. “New technologies are only valuable when they create a profitable business model that helps clients shorten development timelines and secure orders, thereby fostering growth across the entire biotech ecosystem,” he said.
This commitment to value creation resonates deeply with Mycenax’s corporate culture. Dr. Chen explained that since its transition to a CDMO, the company has established a strategic positioning of “Big D and Strong M,” combining robust R&D with high-quality manufacturing. Instead of competing with global CDMO giants on production capacity, Mycenax focuses on advancing emerging biologics such as bispecific (multispecific) antibodies and ADCs. By adopting Cytiva’s advanced technologies, the company demonstrates its capacity to develop challenging novel drugs and optimize low-efficiency processes, supported by a proven track record in GMP-compliant commercial manufacturing.
From ADC to “XYC”: Striking the Balance Between Quality and Capacity
Continuous innovation in biologics design consistently tests the boundaries of downstream bioprocessing. Dr. Chen illustrated this point by using ADCs as an example. Beyond the original combinations of antibodies bound to small-molecule drugs, monoclonal antibodies can be replaced by bispecific antibodies, and the payload can be swapped for nucleic acids, peptides, or radioactive substances. The combinations are endlessly diverse, to the point where they can be termed “XYC” (meaning X and Y have various pairing possibilities). Faced with clients’ endless new concepts and a vast array of drug molecules, the laboratory and facility resources of CDMOs are ultimately limited; it is impossible to infinitely purchase new equipment or constantly modify GMP production lines for every new product.
Against this backdrop, digital twin technology emerges as the optimal solution to address these operational bottlenecks. According to Dr. Chen, the combination of high-performance computing and GoSilico’s mechanistic modeling dramatically broadens the scope of traditional development methods. Whereas classical methods were constrained by high sample costs and time limitations, which restricted testing to just a few scenarios, this platform can run simulations of 3,000 combinations within a single day. This allows process engineers to achieve an optimal balance among compliance standards, product recovery, and yield in early development phases, avoiding the scenario in which a process performs well in a laboratory setting but later proves unworkable in a GMP manufacturing facility.
Grounded in Real Data: Finding the “Golden Batch” with Digital Twins
To fully implement digital twin technology in downstream bioprocessing, substantial volumes of high-quality real-world data remain essential for model calibration, enabling AI to assist users in identifying the “golden batch”, where all process conditions are precisely aligned for optimal quality and yield.
Nevertheless, Jeff Wang acknowledged that achieving these goals remains highly challenging at present. Looking across Taiwan and around the world, many drugmakers have not yet fully adopted electronic records, with some still relying on handwritten paper records or outdated operating systems. Without digitized data, pharma companies are fundamentally unable to determine whether their output is a “golden batch,” making subsequent AI applications and process optimization impossible. The successful implementation of GoSilico at Mycenax serves as a strong proof of concept (POC) for the industry, demonstrating that digitization is not only feasible but also a crucial tool for future competitive edge.
Overcoming Regulatory and Cost Challenges: Accelerating POC Via Platformization
Beyond technological innovations, the biotech industry is also facing a persistent conflict between innovation speed, regulatory compliance, and cost control. Jeff Wang stressed that balancing these forces requires understanding the client’s actual circumstances. For example, in the biosimilars sector, intense price competition has driven the manufacturing costs of monoclonal antibodies down to a range of $100 to $200 per gram. Under this financial strain, equipment suppliers must assist clients in scaling up production by continuously reducing costs and promoting product standardization.
From a regulatory perspective, the biopharma industry operates under strict oversight; the standards set by the FDA and the EMA are non-negotiable, and compromise on compliance for the sake of cost-cutting is never an option. “Innovation speed and production costs are two sides of the same coin,” observed Wang. “Equipment suppliers and CDMO players have long been inextricably linked as a ‘community of shared destiny.’” Therefore, suppliers must offer clients legally compliant reference cases tailored to their current situations, ensuring that R&D and manufacturing solutions scale smoothly without delays.
According to Wang, when clients face stringent inspections from regulatory agencies, they often turn to equipment suppliers first for suitable solutions. Given this reality, suppliers can no longer rely on the traditional business model of merely selling hardware. Instead, they must collaborate with industry partners to understand genuine market needs and deliver customized solutions that satisfy rigorous regulatory standards.
To navigate these regulatory and competitive pressures, Wang emphasized that suppliers must promote equipment “platformization” and “standardization” (such as single-use technology platforms) to accelerate the proof-of-concept stage. For example, the alliance with Mycenax to deploy GoSilico sought to quickly demonstrate the feasibility and cost-effectiveness of the system. He believes Cytiva must continuously validate innovative technologies to maintain its leadership in the competitive global biopharma industry.
Leveraging High-Tech Competitiveness to Lead Globally as a “Boutique CDMO”
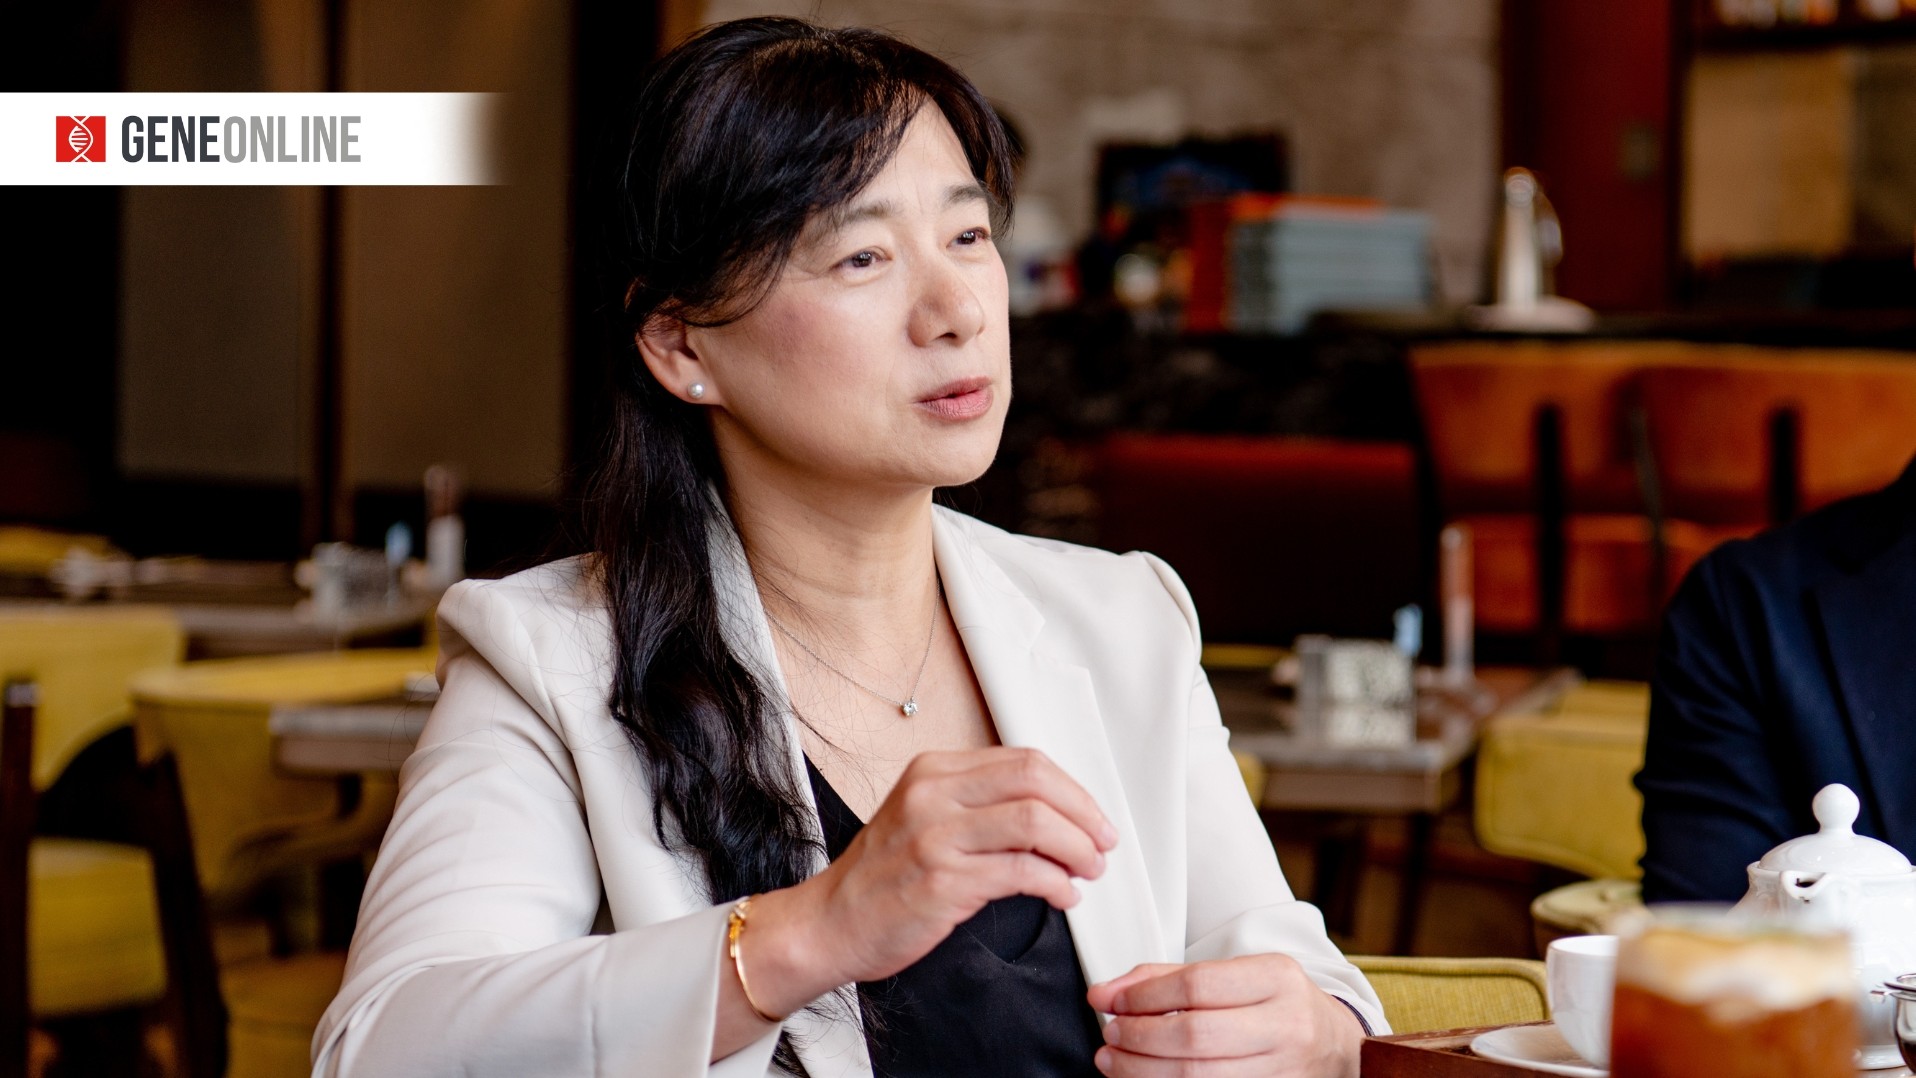
While geopolitical shifts are presenting new challenges for global supply chains, they also create unique opportunities for Taiwan’s biotech industry. Under the leadership of CEO and President Pei-Jiun Chen, Mycenax has successfully passed GMP inspections in multiple countries since 2024, enabling its products to reach 13 markets, including 10 European nations, Canada, Japan, and Korea. In November 2025, Mycenax partnered with three Japanese companies, including Alfresa Holding (one of Japan’s largest prescription drug distributors), Kidswell Bio, and Chiome Bioscience, to establish the joint venture Alfenax Biologics. The company also received a subsidy from the Japanese government to construct a local manufacturing facility.
In the interview, Dr. Chen credited Taiwan’s success to its national competitiveness, including its pivotal geographic location in the APAC region, world-leading AI and ICT technologies, talented and diligent workforce, business regulatory framework aligned with global standards, and strong IP protection, collectively shaping an unparalleled national reputation. Mycenax has effectively leveraged these advantages to establish its strategic blueprint: “rooted in Taiwan, backed by Japan, and advancing into the US and Europe.”
Regarding Mycenax’s operations in Japan, Dr. Chen noted that the Japanese market is relatively reserved, requiring long-term efforts to cultivate connections and build credibility. Nevertheless, once trusted by clients, they demonstrate exceptionally high retention and loyalty. Therefore, the company’s strategy is to retain its cutting-edge technologies and process development capabilities in Taiwan, while building production facilities in Japan to create a dual-base structure, thus enhancing the overall stability and safety of the supply chain.
When asked about the outlook for the CDMO landscape over the next three to five years, Dr. Chen clearly pointed out that Taiwan’s challenge lies in its small market size. It is unrealistic to engage in a capital-intensive battle with global giants boasting production capacities measured in hundreds of thousands of liters. Hence, the focus for future competitive advantage will shift from production scale to the mastery of frontier technologies and digital transformation. She highlighted that Taiwan should position itself to create “Boutique CDMOs,” leveraging technical excellence and high flexibility to resolve complex challenges from clients worldwide. “Our goal is to demonstrate to the global community that Taiwan has a robust biomanufacturing sector alongside its world-class semiconductor industry, and we need to strive to cultivate trust among international developers toward Taiwan’s CDMO enterprises,” remarked Dr. Chen.
Going Global by Trusting Professionals and Keeping the End in Mind
Concluding the interview, both executives drew upon their extensive industry experiences to offer valuable suggestions for early-stage biotech startups. Dr. Pei-Jiun Chen observed that start-up teams originating from hospitals, universities, or research institutes often place excessive emphasis on initial drug efficacy while overlooking the numerous considerations involved in the commercialization process. She emphasized that regardless of how promising these laboratory findings might appear, they become impractical if they cannot be smoothly transitioned into large-scale, GMP-compliant manufacturing.
Therefore, she advised these start-ups to adopt a proactive approach by “beginning with the end in mind”. This involves integrating future commercial-scale production requirements into their planning from the very inception of an R&D project, while also engaging in early, detailed discussions with CDMO partners with advanced equipment and specialized expertise. Taking these steps ensures that their hard-earned R&D breakthroughs remain on the right track, effectively preventing devastating setbacks—such as having to scrap and restart the entire project—when scaling up production later on.
Jeff Wang added that many new companies, after securing funding, often think first of investing heavily in building their own manufacturing facilities, but he stated bluntly that this is actually a massive misallocation of resources. He emphasized, “A company’s core value lies in intellectual property (IP), not in its manufacturing facilities. Rather than spending vast sums on maintaining facilities and navigating related compliance mandates, it is far better to prioritize resources toward demonstrating the drug’s efficacy.”
Wang also addressed start-ups’ reluctance to entrust their efforts to third parties, stressing that trust is key to overcoming this mentality. He argued that in the modern business landscape, it is essential to move past the outdated mindset of working in isolation or handling everything in-house. Instead, delegating manufacturing to professional CDMO teams is the most efficient way to increase success rates and accelerate time-to-market.
Furthermore, highlighting Taiwan’s world-class medical professionals, reliable National Health Insurance database, and regional biotech clusters, Wang urged biotech start-ups to leverage this local ecosystem to focus core resources on clinical proof-of-concept studies. Through flexible resource integration, trust in partners, and a clear division of labor, he believes Taiwan’s biomedical industry will be able to “grow the pie” and compete on the global stage.
Agile Manufacturing and Process Intensification: Single-Use Technology and Continuous Purification
The “Building Future-Ready Downstream Bioprocessing” seminar featured 7 expert presentations. The morning sessions explored process intensification through single-use platforms, continuous processing, and PUPSIT regulatory practices, guiding attendees on designing flexible, compliant, and scalable biomanufacturing lines.
Dr. Albert Liao, Head of Manufacturing Science and Technology (MSAT) at Mycenax, highlighted the strategic value of single-use platforms. He noted that rising global demand for biopharma CDMO outsourcing brings challenges such as multi-product concurrency, diverse batch sizes (ranging from small early-stage clinical lots to commercial mass production), tight R&D schedules, and cross-contamination risks. As a result, traditional stainless-steel equipment lacks the flexibility needed for the modern biotech industry.
Dr. Liao explained that stainless-steel systems entail high upfront costs and work best for long-term mass production of a single product. However, multi-product concurrent runs trigger complications, forcing facilities to halt operations for thorough cleaning, swab sampling, and repetitive worst-case evaluations. Remaining residues from incomplete cleaning are classified as major GMP deficiencies by regulatory bodies. Conversely, single-use systems leverage disposable components such as storage bags, tubing assemblies, and filters, eliminating tedious cleaning and sterilization. This cuts water usage by nearly 90%, reduces energy consumption by 50-60%, and drastically shortens batch changeover times, enabling CDMO facilities to switch between products rapidly and boost operational adaptability.
Dr. Ming Yue Fang, Head of China NPP & BRD Digital Intelligence Innovation at WuXi Biologics, gave an online talk titled “Downstream Process Intensification: From Batch to Continuous Purification”. Leveraging her extensive background in managing over a hundred cell culture and process development initiatives and establishing WuXi Biologics’ ultra-high productivity continuous bioprocessing platform, WuXiUP™, she detailed the numerous benefits of continuous processing.
In the development of antibody biologics, breakthroughs in upstream cell culture and fermentation have led to high-concentration, high-viscosity feedstocks, creating bottlenecks for traditional batch purification due to equipment and time constraints. In contrast, continuous purification maximizes efficiency and output within smaller facility spaces. To address this shift, Dr. Fang shared case studies and process intensification strategies to help biotech companies successfully transition from conventional batch processing to continuous purification.
Navigating the Regulatory Expectations for PUPSIT Implementation
Patient safety relies heavily on the strict sterility control of biologic drugs, serving as an uncompromising regulatory baseline. In the fourth session of the seminar, Inna Snisarenko, Cytiva’s Global Product Manager, explored this complex compliance hurdle in her presentation, “Regulatory expectations and practical solutions for PUPSIT implementation”. She underscored the significant changes in the EU GMP Annex 1 guidelines, noting that the revised 2022 version expanded significantly to 58 pages from the mere 16 pages comprising the 2008 edition.
The updated 2022 guidelines explicitly mandate incorporating quality risk management (QRM) into the contamination control strategy (CCS). Also, manufacturers must perform Pre-Use Post-Sterilization Integrity Testing (PUPSIT) to verify that filtration systems remain undamaged during preparation and sterilization, thereby blocking potential sterility breaches. This regulatory shift impacts not only European markets, but also all countries adopting the PIC/S GMP guide—such as Taiwan, Singapore, and Korea—as well as any global manufacturer exporting sterile drugs to Europe.
Traditional PUPSIT is complex and prone to leaks, dilution, or product adsorption during manual filter wetting, pressure testing, and venting. To mitigate these bottlenecks, Cytiva offers automated, standardized PUPSIT solutions by integrating single-use flow kits with automated drug product filtration systems to minimize human error. These advanced filtration units feature automated leak and integrity testing, while recipe-driven configurations guarantee batch uniformity, aligning with European contamination prevention mandates to balance efficiency and compliance.
Overcoming the High-Concentration Biologic Challenges
To reduce medical expenses and enhance patient quality of life, pharmaceutical companies have increasingly adopted subcutaneous delivery methods for biologics in recent years. Yet, such dosage forms typically require elevated drug concentrations reaching 100 g/L or even 200 g/L, introducing serious hurdles for downstream purification workflows. During the seminar’s fifth session, Wang Jing, Cytiva’s Manager of Filtration Modality, explained that high-concentration feeds exhibit increased viscosity and a higher tendency to generate protein aggregates. These characteristics frequently trigger a rapid decline in filtration flux and premature filter blockage, ultimately diminishing product recovery. To address the purification challenges associated with these highly concentrated biologics, Wang outlined two key technological solutions:
- Single-pass tangential flow filtration (SPTFF): Compared to traditional TFF, SPTFF eliminates the conventional recirculation tank, which not only reduces the system’s working volume but also vastly decreases the number of pump passes and shear exposure. This technology can effectively mitigate the risk of protein aggregation; in experiments, it successfully concentrated monoclonal antibodies to over 200 g/L with recovery rates as high as 95%.
- Supor™ Prime 0.2 µm sterilizing grade filter: Featuring an innovative asymmetric and symmetric dual-layer PES membrane architecture, this novel filter achieves excellent flux rates when processing high-concentration antibody feedstocks with diverse particle sizes. Thanks to its outstanding filtering performance, facilities can use smaller-sized filter cartridges, effectively reducing the hold-up volume within the assembly. This design also reduces flush volume requirements and minimizes product loss due to adsorption, resulting in a substantial increase in overall batch yield.
From Data Integrity to High-Efficiency GMP Operations
Digital operations are critical for facility efficiency. In the sixth presentation, Sharon Huang, Account Manager at Rockwell Automation, outlined a blueprint for GMP plants to move from data integrity to operational excellence. She noted that many biopharma plants still rely on manual operations and paper records, while others face “system silos” where R&D and manufacturing data are disconnected, and base-level sensor data fails to integrate into the enterprise resource layer. This hampers the replication of manufacturing knowledge across facilities. To overcome these challenges, she urged the industry to adopt digital platforms. For instance, implementing Rockwell’s Electronic Batch Recording (EBR) eliminates tedious paper-based tasks, reduces manual entry errors, and significantly shortens batch-review cycles to accelerate product releases.
Huang also emphasized the massive potential of combining industrial data operations with AI. Integrating data from base-level equipment, information systems, and engineering designs into a contextualized industrial knowledge graph allows AI to understand complex factory data. Consequently, future factories will gradually shift from passive, descriptive models to AI-driven predictive and prescriptive systems, ultimately achieving autonomous operations with auto-debugging and continuous optimization capabilities.
Tackling CMC Submission and Compliance Issues
Dr. Ang Wei Xia, Senior Regulatory Compliance Lead at Cytiva, delivered the seminar’s final presentation on quality considerations in manufacturing for antibody CMC submissions. Drawing from her background at Singapore’s Health Sciences Authority (HSA) and Agency for Science, Technology and Research (A*STAR), she explained that chemistry, manufacturing, and controls (CMC) define production, characterization, and release criteria for drug substances and products to ensure safety and consistency. For complex biologics like monoclonal antibodies, regulatory bodies heavily scrutinize structural characterization, impurity control, process robustness, and traceability. Consequently, establishing and validating effective downstream purification steps is critical to guaranteeing final product safety and purity.
Addressing key sterile filtration compliance challenges, Dr. Ang examined the 2022 revised EU GMP Annex 1 guidelines on PUPSIT practices. She explained that the new requirement mandating PUPSIT prevents “flaw masking”, where a filter damaged during sterilization allows bacteria to penetrate but is later plugged by product debris, causing a false “pass” during post-use integrity testing. To meet these regulations while maintaining operational safety and product yield, Dr. Ang proposed the following practical recommendations:
- Keep it safe and simple (KISS): Prioritize patient safety by implementing quality risk management (QRM) to prevent convoluted pipeline designs that could elevate risks of contamination.
- Consider redundant filtration: Configure dual sterile filters as a compliant backup in case the primary filter fails the post-use integrity test, preventing the disposal of an entire batch of precious drug liquid.
- Embrace automated filtration systems: Adopt standardized single-use flow kits and automated filtration systems. Their integrated leak and integrity testing reduce manual intervention and errors, harmonizing productivity with rigorous regulatory standards.
Dr. Ang concluded her presentation by highlighting Cytiva’s specialized regulatory consultation services. Supported by a network of over 100 experts, Cytiva can guide biotech companies through compiling CMC submission dossiers, preparing for FDA or EMA site inspections, and developing customized regulatory roadmaps for each phase of drug development, thereby accelerating the timeline for global market entry.
Author
LATEST
EVENT
2026-10-20
AusBiotech 2026 (20-23 Oct, Queensland) (Australia biotech & investment)
Queensland, Australia